Multiscale kinetic modeling in catalysis ⇒ from microkinetics to computational fluid dynamics and process simulations


Multiscale modeling (MKM)

Multiscale kinetic modeling in catalysis: from microkinetics to computational fluid dynamics and process simulation. We connect models from the elementary step to the plant. Each scale passes parameters to the next, and design questions travel back down. Models are built together with the experiments of the other three competencies and are validated against them, so experiments are planned with the model, not guessed.

Problem statement. Catalytic processes occur at several scales at once: elementary steps on active sites, diffusion and heat transfer inside particles, hydrodynamics in the reactor and integration in the process. A model at one scale is not enough when the limiting phenomenon sits at another. Kinetic models fitted to limited or noisy data are not robust enough for scale-up.

We see multiscale modeling as the tool that links reaction understanding, catalyst and reactor design, scale-up and process optimization. We address each scale individually and the chain as a whole.

Our approach, scale by scale. The video zooms from the plant to the electrons, and each paragraph below matches one scene.

1. Process: the plant as a whole. Process models evaluate recycles, heat integration, energy use and scale-up scenarios. We use process system engineering (gPROMS and other packages) for ammonia crackers and reformers, including repurposing methane steam reformers as ammonia crackers [175], and for hydrocracking units, including techno-economic analysis [148]. These models need a reliable reactor model inside.

2. Reactor: flow, heat and solids. We couple hydrodynamics, heat transfer and kinetics in computational fluid dynamics:

  • CFD (Ansys Fluent) for gas-solid packed beds and operando cells [124] ;
  • CPFD (Barracuda) for fluidized beds, downers and risers, validated with particle image velocimetry [162, 181, 184] ;
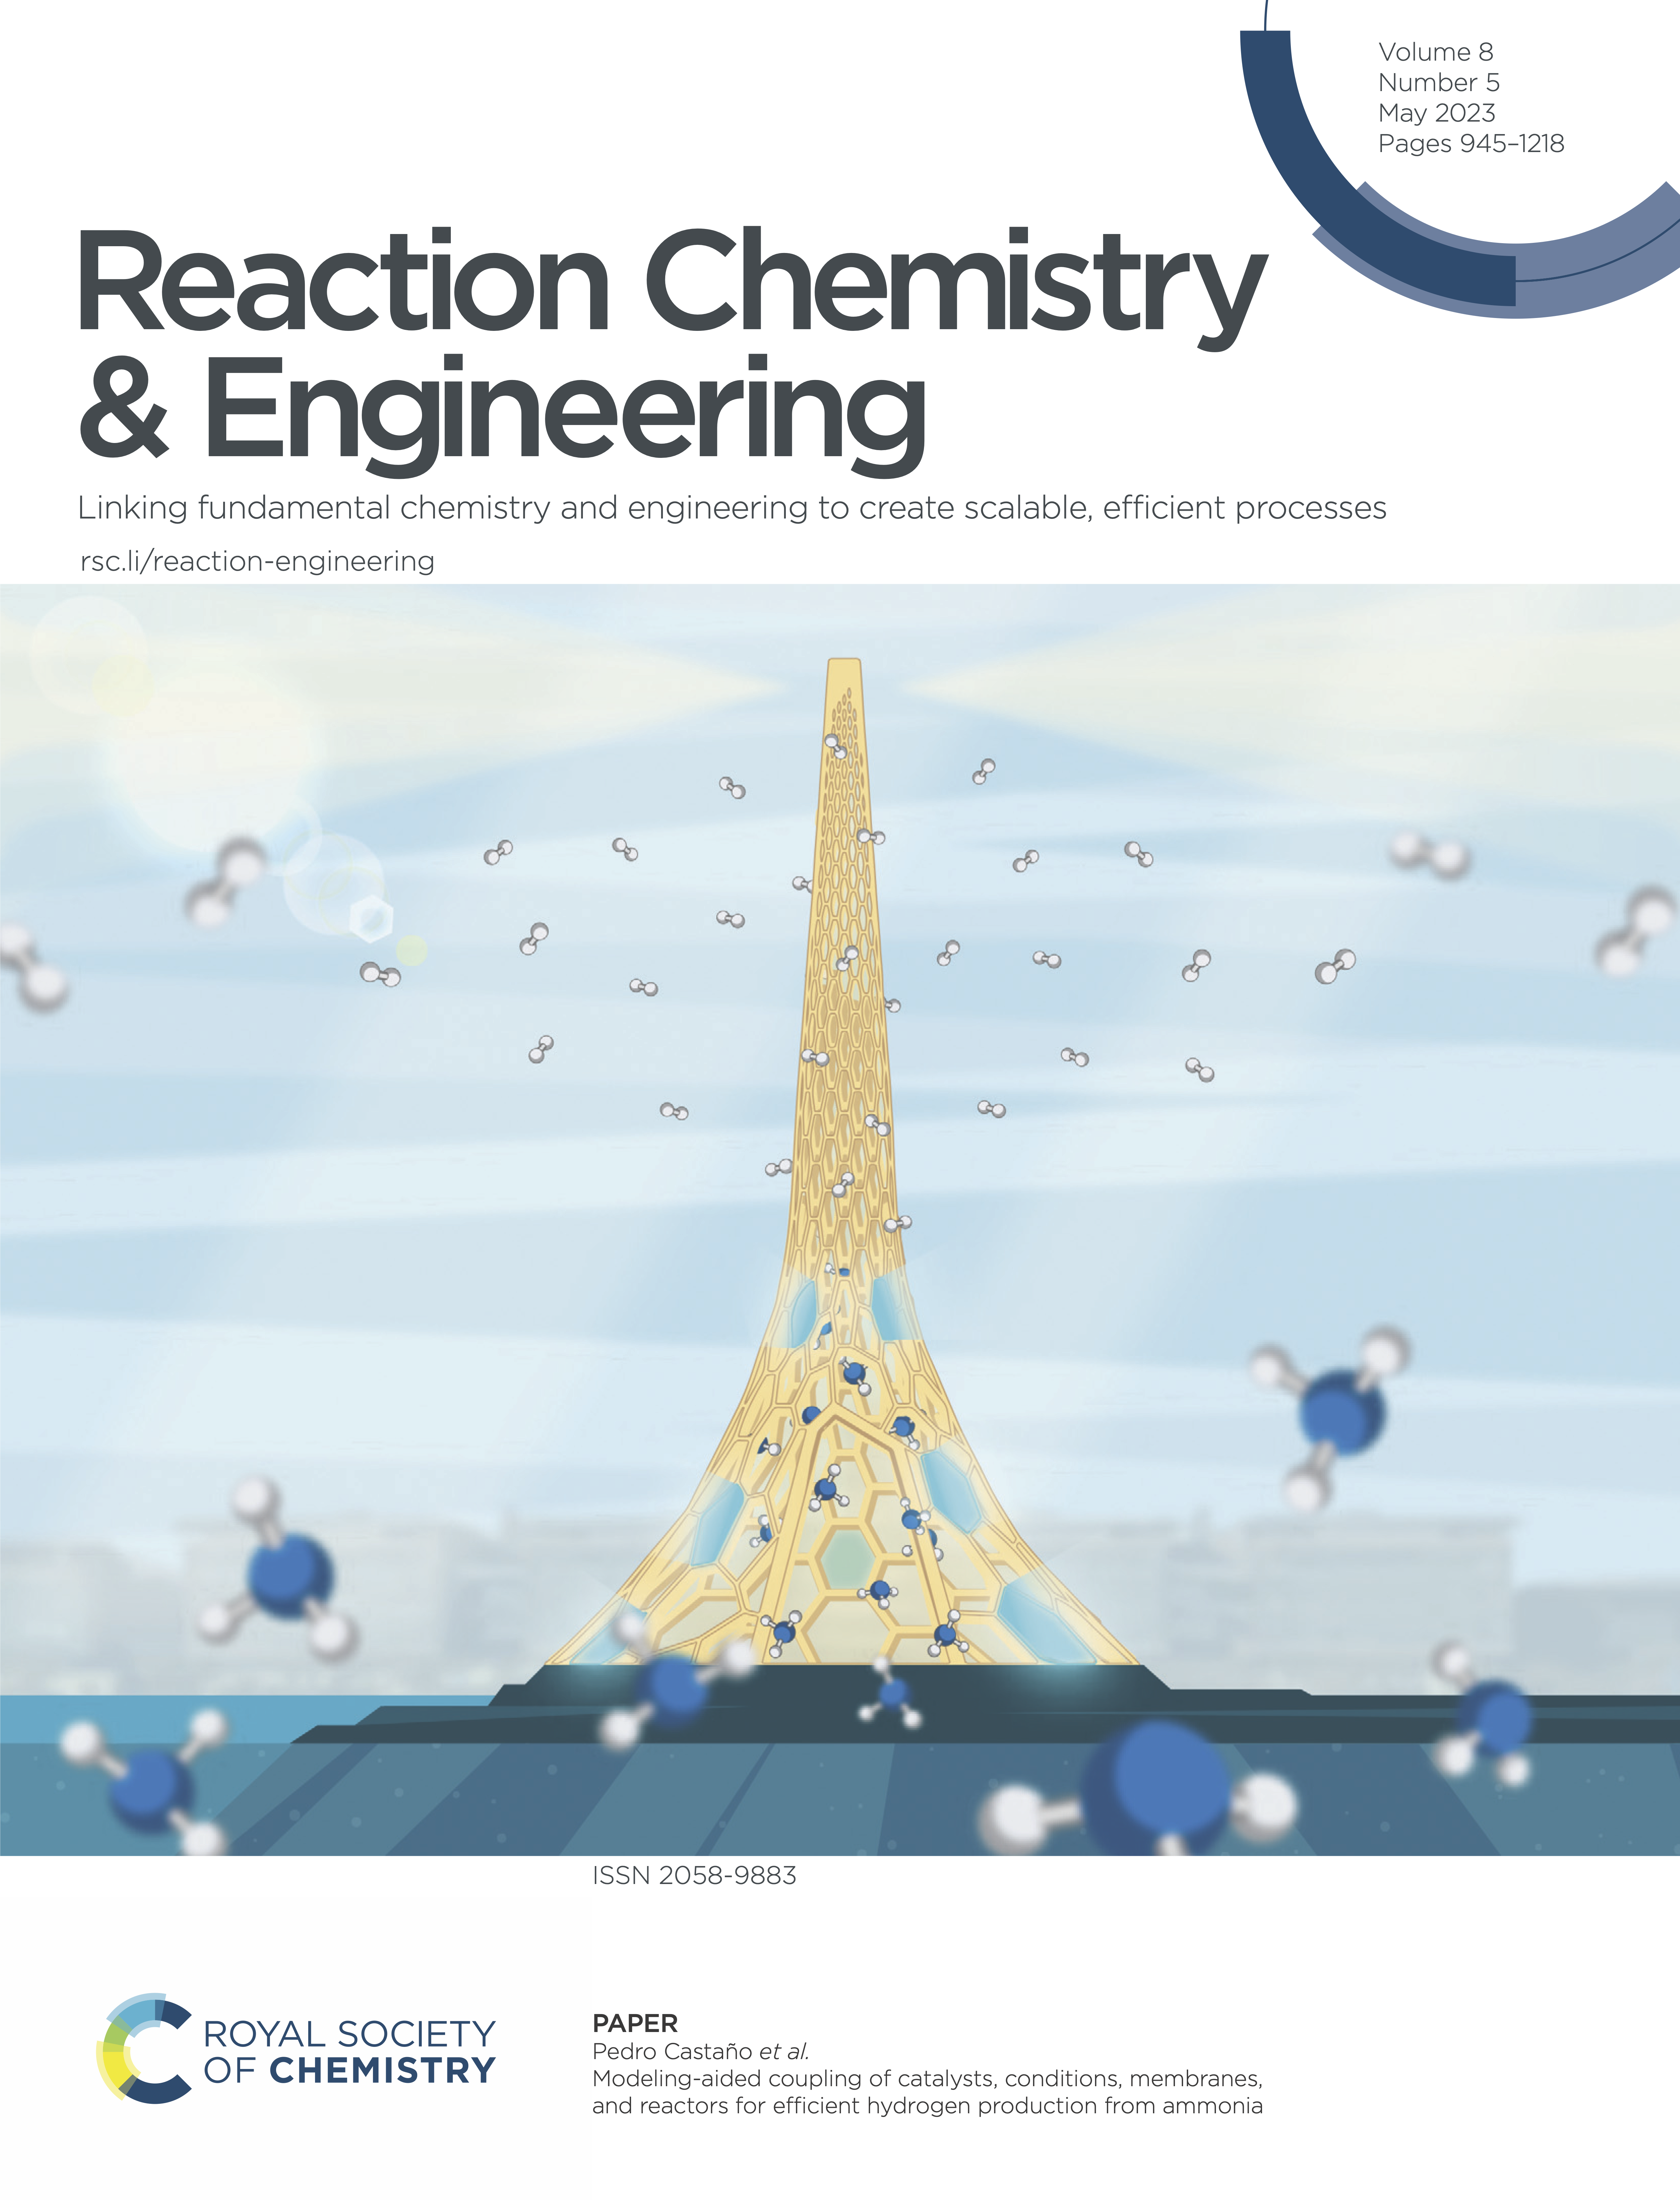
  • dimensionless and reactor models for membrane reactors producing pressurized, high-purity H2 from ammonia [116, 142] .

3. Particle: reaction against transport. Inside each particle, reaction competes with diffusion and heat transfer. This determines how much of the catalyst actually works. Particle models connect intrinsic kinetics with the shaped catalyst, including the effect of SiC and of the shaping method [190], and population balances describe how particles grow during shaping [204].

For complex feeds, macrokinetic models (lumped and empirical) are often the most robust choice. They detect changes in mechanism, optimize conditions and support reactor pre-design. We make them robust with:

  • lumping and continuous lumping strategies for crude-to-chemicals cracking and plastic-oil hydrocracking [177, 195, 206] ;
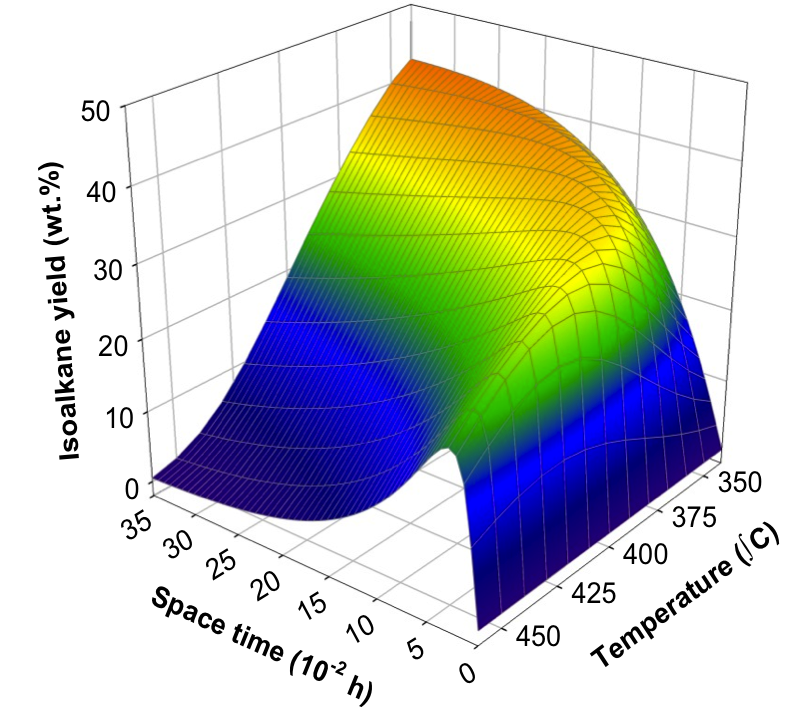
  • acidity descriptors and lumped models for methanol to hydrocarbons and aromatics [156, 176] ;
  • neural networks combined with DFT for reforming kinetics [117] ;
  • outlier detection and data curation for large datasets from high-throughput reactors [154] .

4. Surface: microkinetics. Microkinetic models resolve the rate of each elementary step, the rate-determining step, and adsorption and desorption, with thermodynamic constraints imposed [168] . Langmuir-Hinshelwood and Eley-Rideal formulations are used where they apply. We applied this approach to promoters and active sites in:

  • Ga on Cu for CO2 hydrogenation to methanol [127, 191] ;
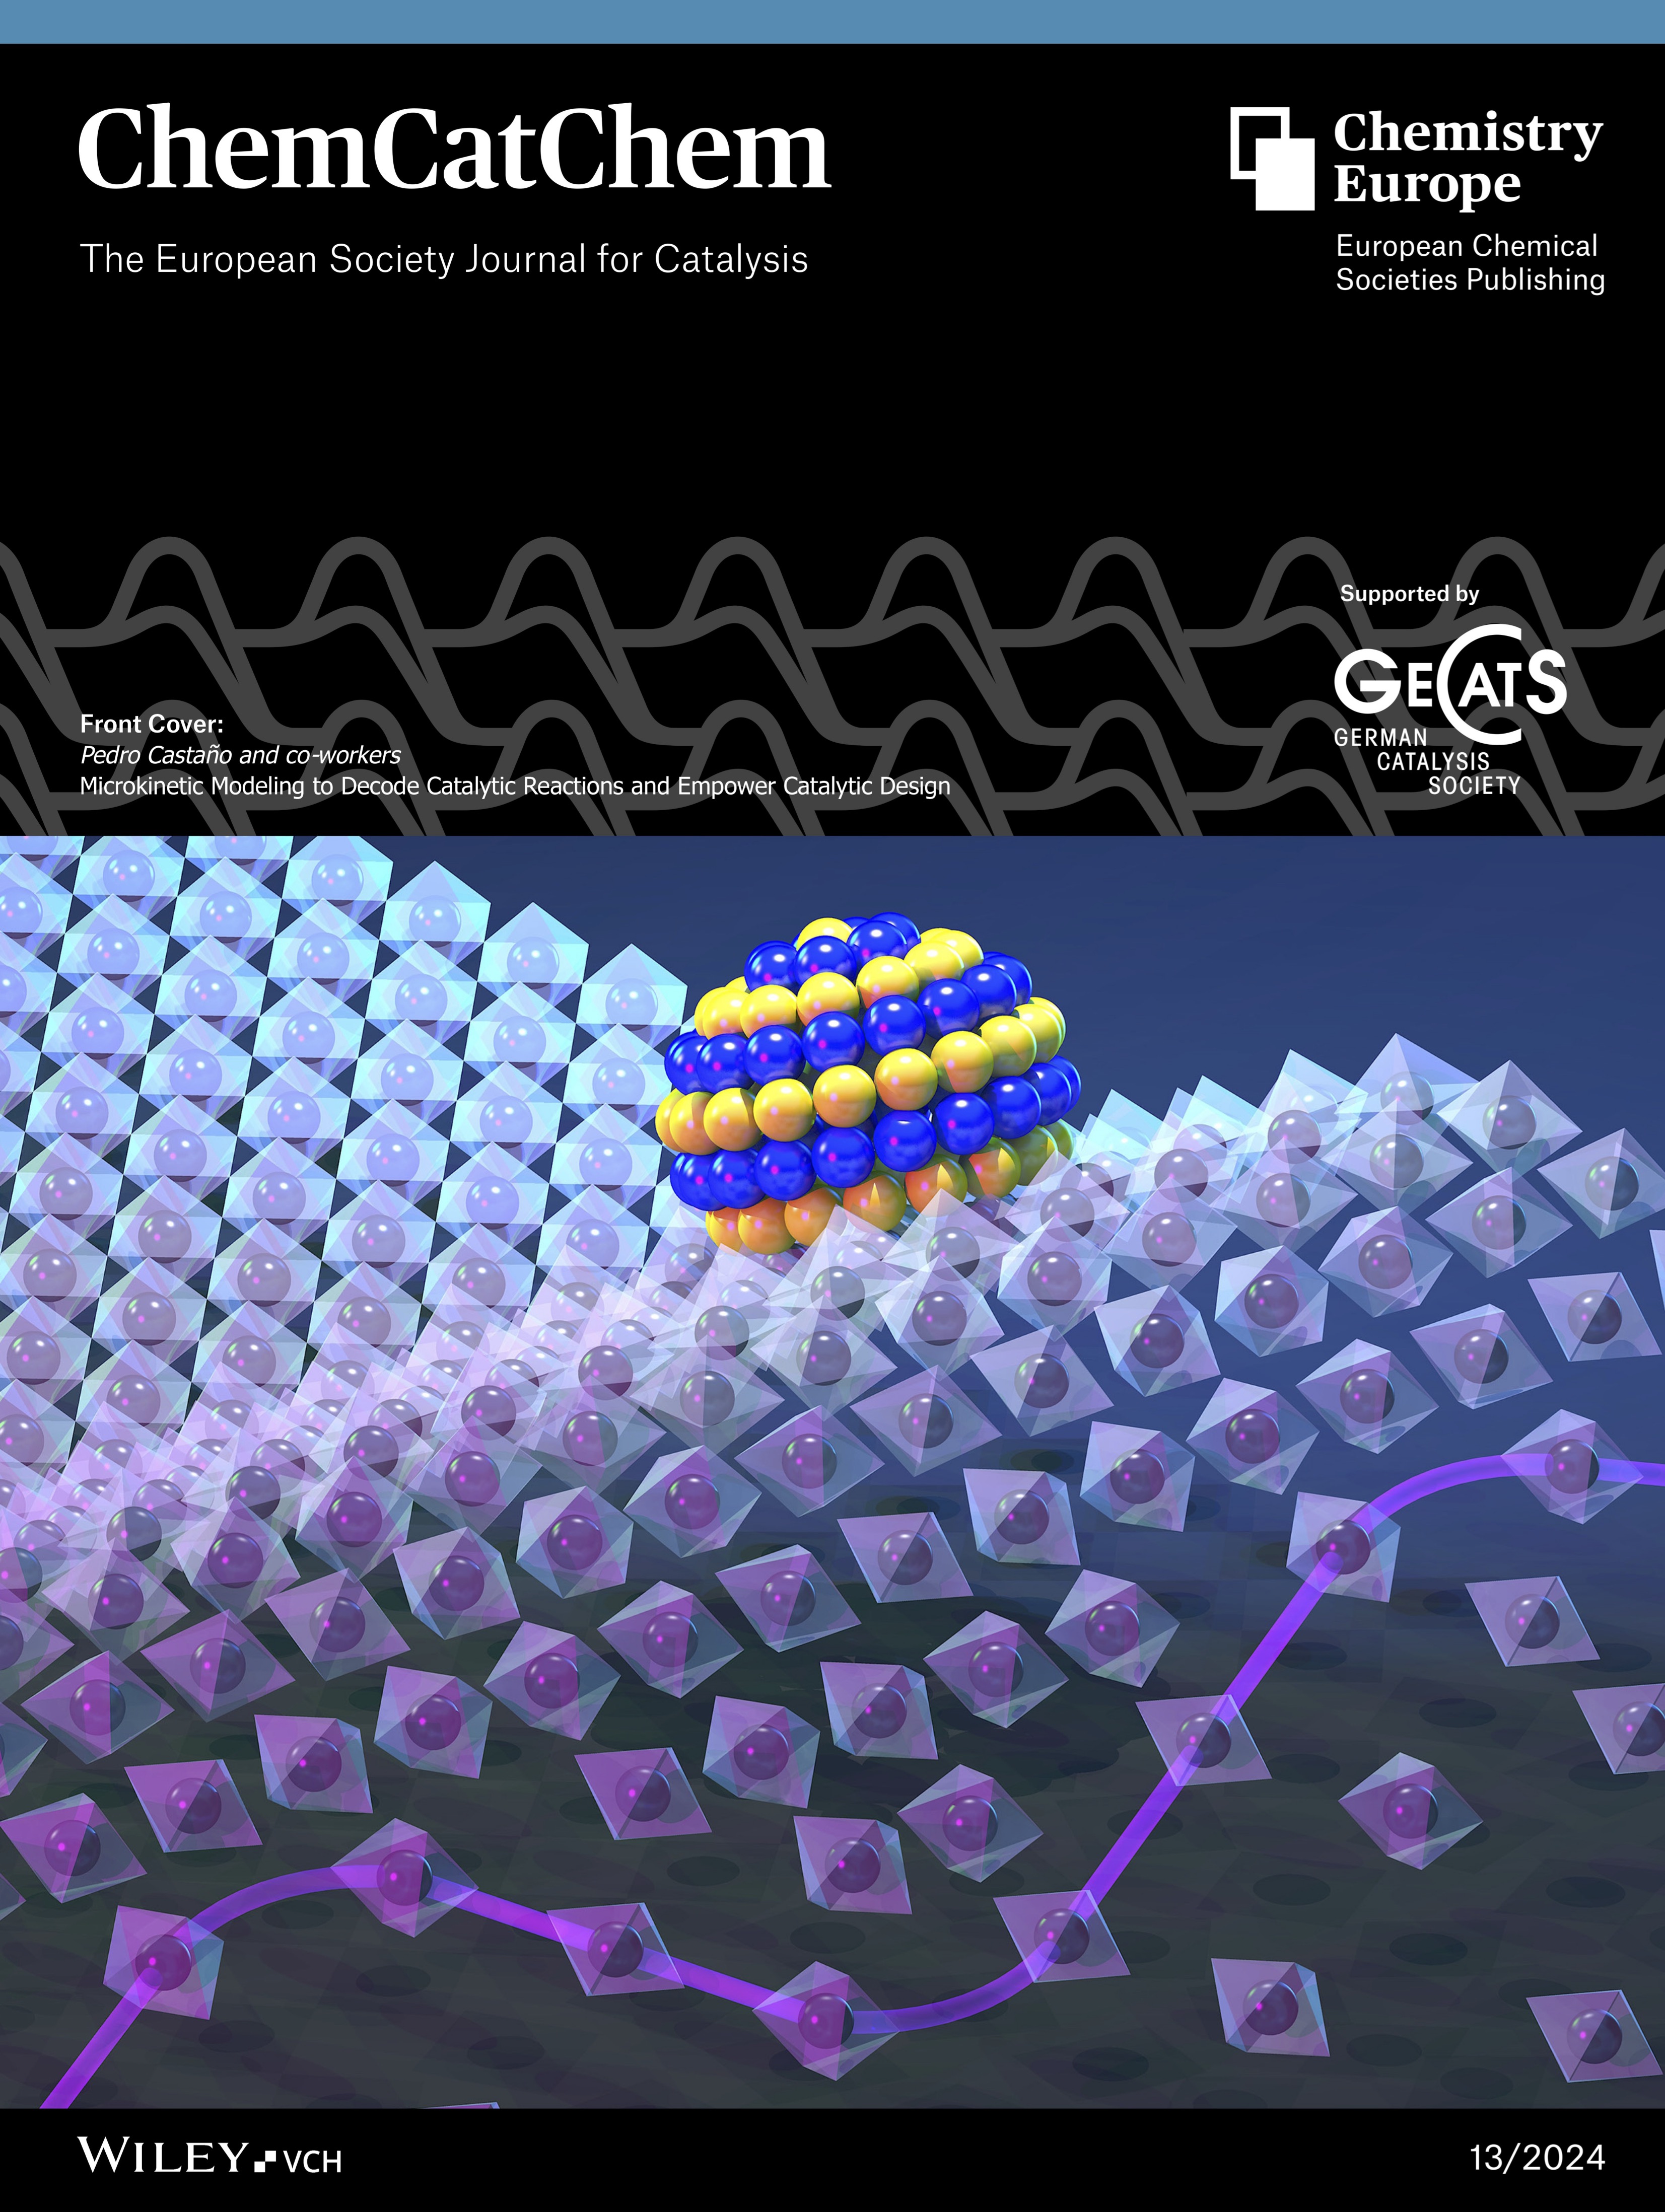
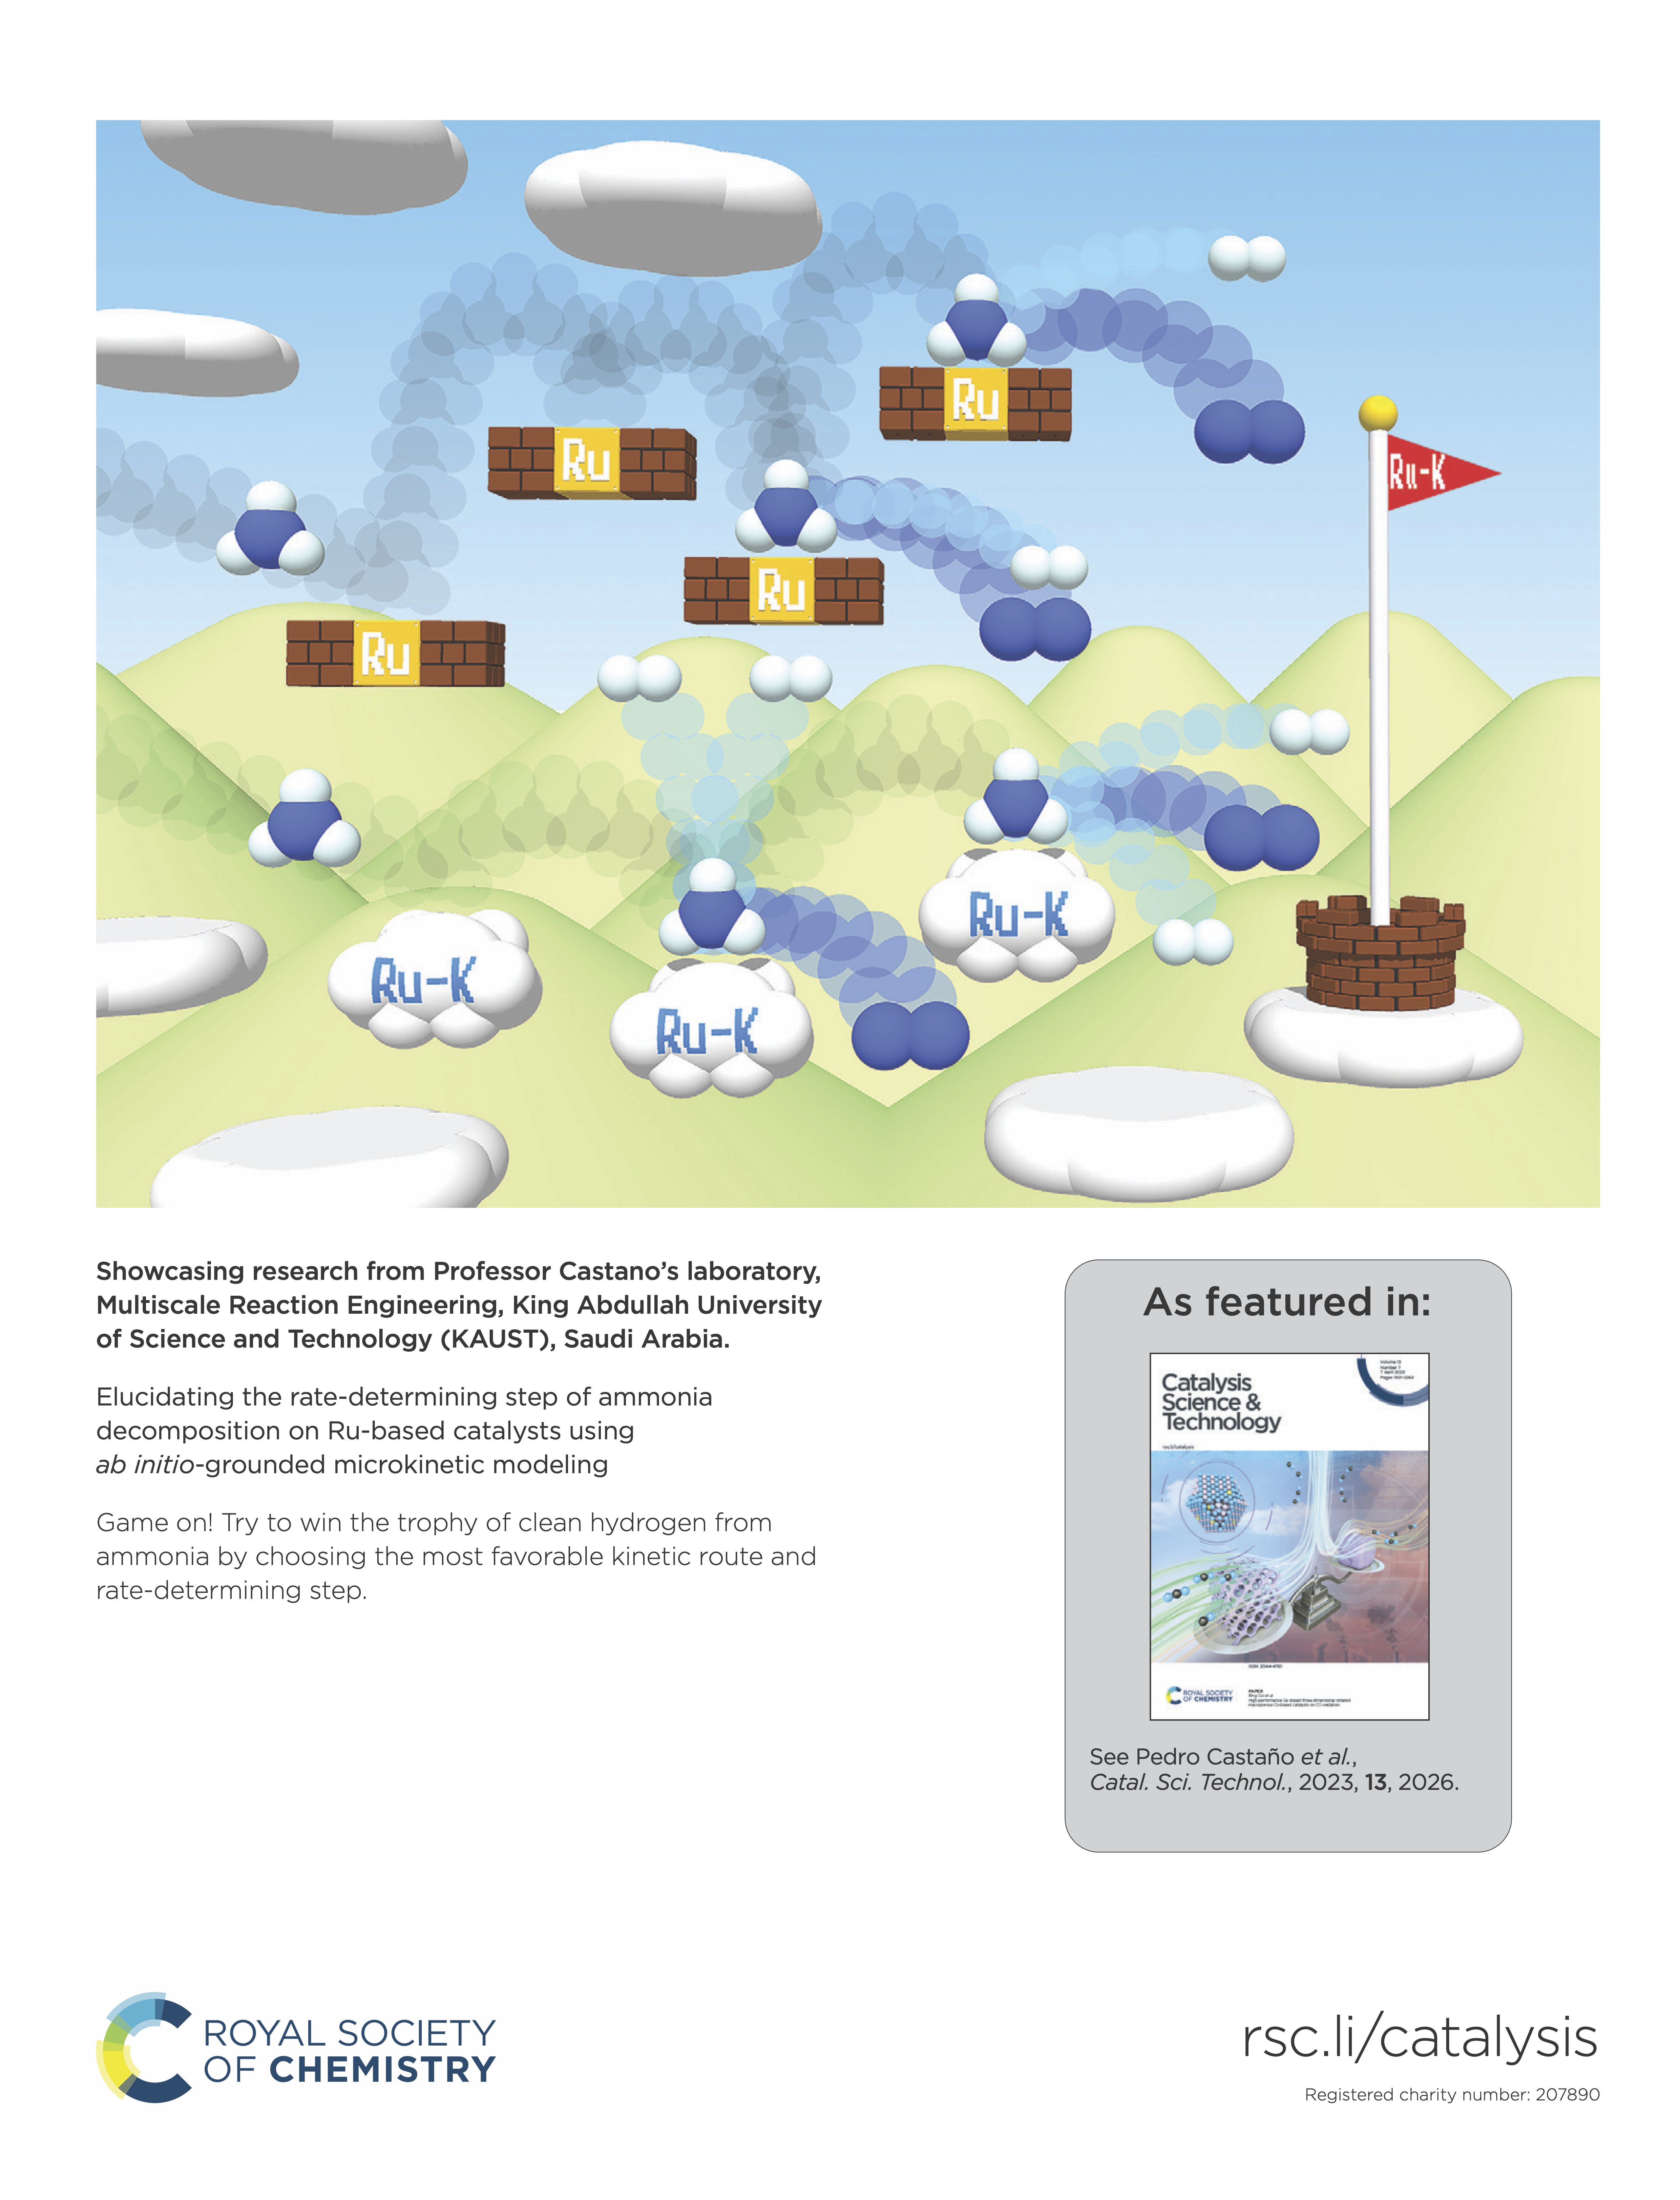
  • K on Ru and Ba on Co-Ce for ammonia decomposition [140, 147] ;
  • microscopic SiC on Mn-Na2WO4/SiO2 for oxidative coupling of methane [190] ;
  • Ni in hollow ZSM-5 for ethylene oligomerization [198] ;
  • sulfide catalysts for H2S dissociation, using reaction descriptors [180] .

5. Electrons: DFT. Density functional theory (DFT) supplies adsorption and barrier energies, supports the assumed pathways and provides initial parameter estimates [140, 180] . From here, parameters move up the chain to the process, and design questions move back down.

Goals. Our goals are grouped in these lines of work:

  • Microkinetics I: key thermodynamic relationships and thermodynamic consistency.
  • Microkinetics II: fitting, training and optimization against kinetic and spectroscopic data.
  • Microkinetics III: ab initio kinetic modeling with DFT energies and reaction descriptors.
  • Macrokinetics: complex reaction networks and population balances.
  • CPFD: fluidized reactor modeling and scale-up.
  • CFD: reactor modeling and optimization.
  • CFD II: modeling operando reactors, to separate kinetics from transport and hydrodynamics.
  • Process system engineering: gPROMS models for scale-up, intensification and techno-economic evaluation.

Applications. We apply this competency to:
  • Ammonia cracking and crude-to-chemicals with Saudi Aramco: multiscale kinetic models guided scale-up to pilot plant in both projects.
  • CO2 hydrogenation, reforming and oxidative coupling of methane.
  • Hydrocracking of refractory, renewable and waste streams.
  • Design of new fluidized and operando reactors, together with the reactor engineering team.

Related People

Related Covers

Related Publications

Comprehensive Approach for Designing Different Configurations of Isothermal Reactors with Fast Catalyst Deactivation

by Cordero-Lanzac, Aguayo, Gayubo, Castaño, Bilbao
Chem. Eng. J. Year: 2020

Abstract

A methodology for simulating the performance of different reactor configurations for processes with complex reaction networks and fast catalyst deactivation has been proposed. These reaction configurations are: packed bed, moving bed and fluidized bed reactors with and without catalyst circulation. From kinetic parameters collected in a packed bed reactor and a rigorous consideration of the activity, modifications in the convection-dispersion-reaction equation have led to the prediction of the catalyst performance in each reactor configuration. The circulating fluidized bed reactor has been simulated with an original model of parallel compartments, which allows for determining its performance in the steady state from the evolution of the transitory period. The methodology has been used for simulating the dynamics of SAPO-34 fast deactivation during the methanol-to-olefins (MTO) process. For each reactor configuration, concentration profiles and their evolution with time have been simulated, thus predicting the effect of reaction conditions and water content (formed and/or co-fed) on the activity profile or the activity distribution function (in the case of circulating fluidized bed reactor). The olefin yield and distribution have also been compared for each reactor configuration.

Keywords

O2H FCC CRE MKM